01
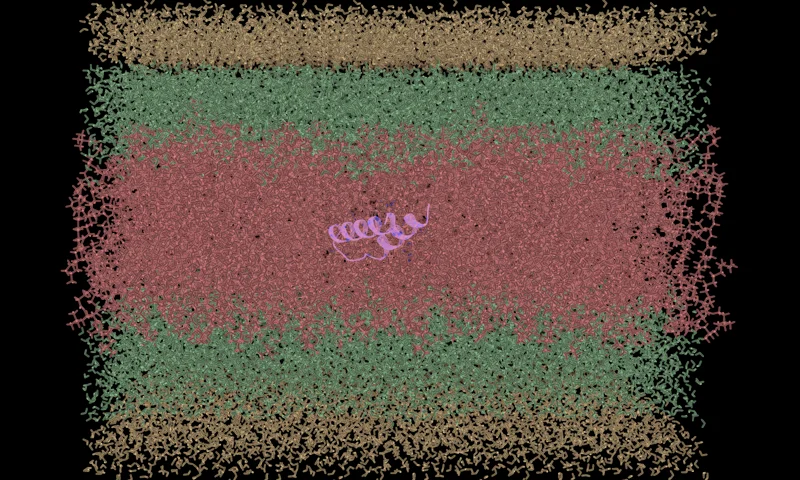
Prepare and relax the system
Repair the starting structure, choose molecular states and force-field assumptions, solvate, minimize, and equilibrate before production sampling.
Molecular dynamics & free energy
Prepare and relax systems, run all-atom or coarse-grained simulations, analyze trajectories, and compare binding energetics without maintaining an HPC stack.
Trusted by researchers at leading biotech, pharmaceutical, academic, and materials organizations


Structure to dynamics
Make force fields, protonation, solvent, boundaries, equilibration, and sampling choices explicit so every trajectory remains interpretable.
Repair the starting structure, choose molecular states and force-field assumptions, solvate, minimize, and equilibrate before production sampling.
Select all-atom or coarse-grained representation, define ensemble and duration, and retain the settings needed to reproduce the trajectory.
Quantify structural stability, interactions, solvent behavior, and relative binding evidence across trajectories or compound series.
Designed around the research
Available now
Preserve setup and sampling choices, assess convergence, and interpret energetic estimates within the scope of the selected method.
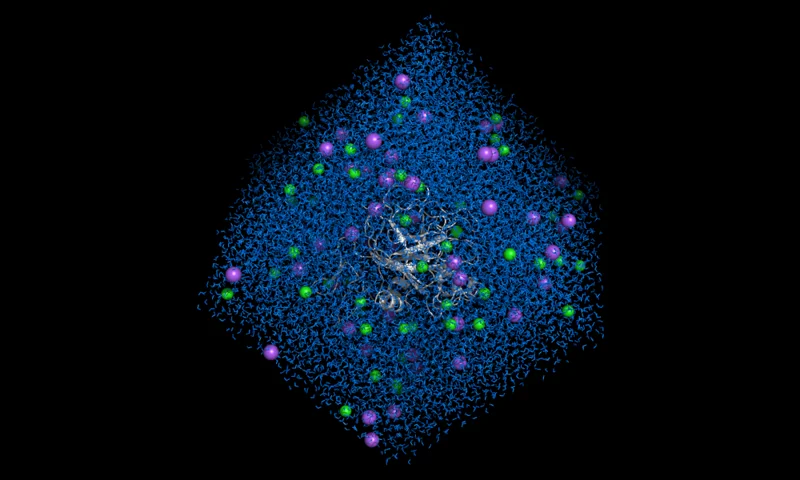
Run production biomolecular simulations.
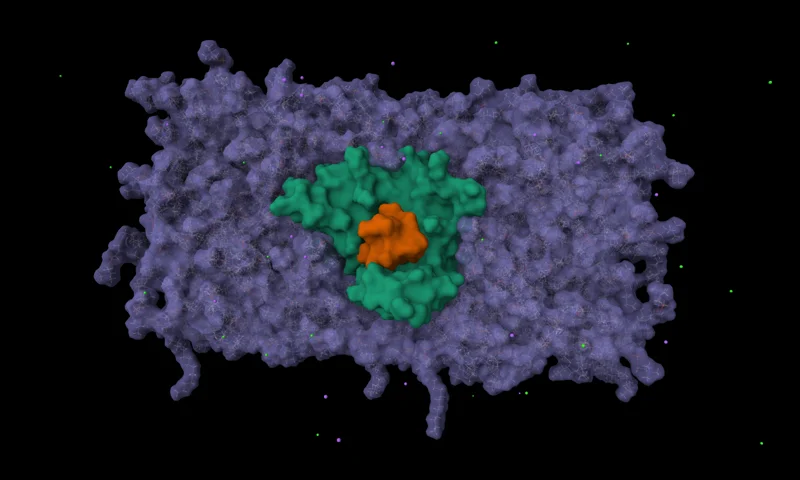
Build and simulate flexible molecular systems.
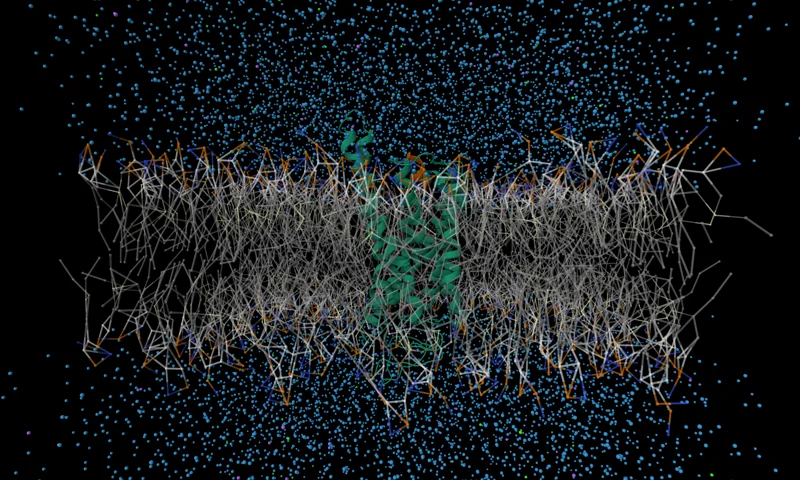
Explore longer scales with coarse-grained dynamics.
Analyze structural and interaction changes over time.
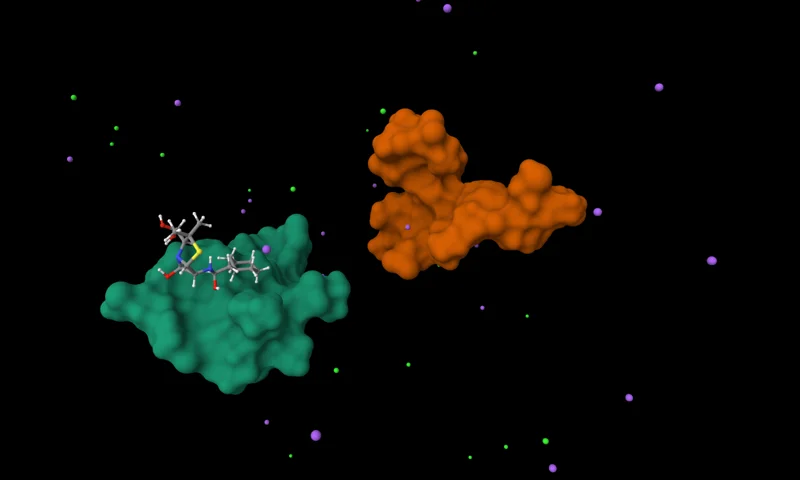
Estimate binding energies from GROMACS trajectories.
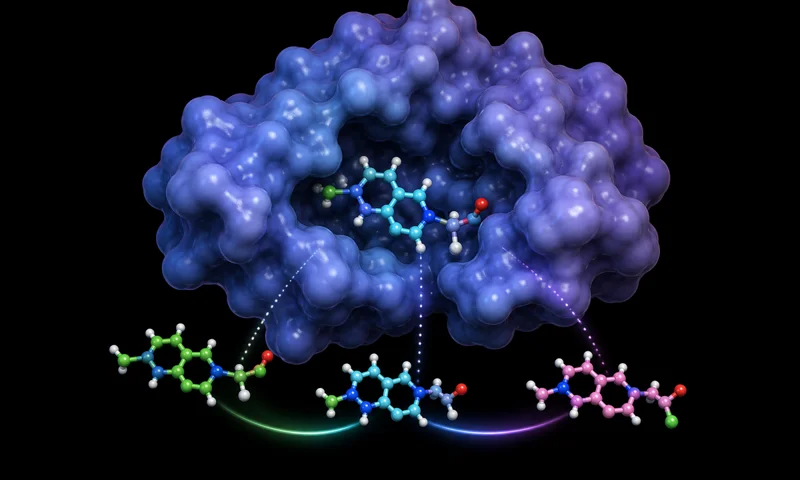
Compare ligands with relative free-energy calculations.
From the Neurosnap blog
Planning guide
See how system size, representation, duration, and preparation choices influence computational cost.
Read articleSimulation guide
Review system setup, equilibration, production settings, and the outputs needed for trajectory analysis.
Read articleSecurity & privacy
Submitted data and generated results remain confidential, with ownership retained by your team.
Only you and authorized colleagues can access submitted data and results.
Your results and intellectual property remain yours. No royalties or equity.
Platform controls help protect sensitive computational research.
Onboarding and integration support help organizations scale confidently.
Sample molecular behavior
Run reproducible simulations, inspect motion and interactions, and carry dynamic evidence into the next research decision.